
Since the previous research note · Earlier expectation not tested
Track continuity
Since the previous research note: How a model follows lithium through a solid electrolyte to the cell's operating limit
Previous expectation: Validated transport models could help researchers match electrolytes to electrodes and current demand, potentially extending thin-film cell operation before cathode saturation.
What this study adds: This source develops the electrolyte description through two transport mechanisms, an energy-aware formulation intended to connect scales, and analysis identifying the equilibrium mobile-lithium fraction as especially influential. These contributions refine the modeling foundation but do not test the earlier forecast of improved cell operation.
What changed in this study
A solid-state battery electrolyte can contain a large amount of lithium without making all of that lithium equally available for transport. That distinction matters because an electrolyte’s modeled response depends not only on how much lithium is present, but also on which ions can move, how they move, and how the surrounding vacancies change as the system evolves.
A July 6, 2021 preprint proposes a theoretical and numerical description built around that problem. Its aim is a three-dimensional model of a solid electrolyte that can represent two forms of lithium-ion motion while remaining compatible with descriptions at different scales. The work studies both equilibrium and steady-state behavior, then examines which material parameters most strongly affect the model’s predictions.
This is a contribution to the machinery used to reason about solid electrolytes. It is not a demonstrated improvement in battery runtime, charging speed, energy density, or safety. Its value lies in making the transport calculation more explicit: lithium can be present in the material, yet only part of it may occupy a modeled mobile state.
Lithium can move in more than one modeled way
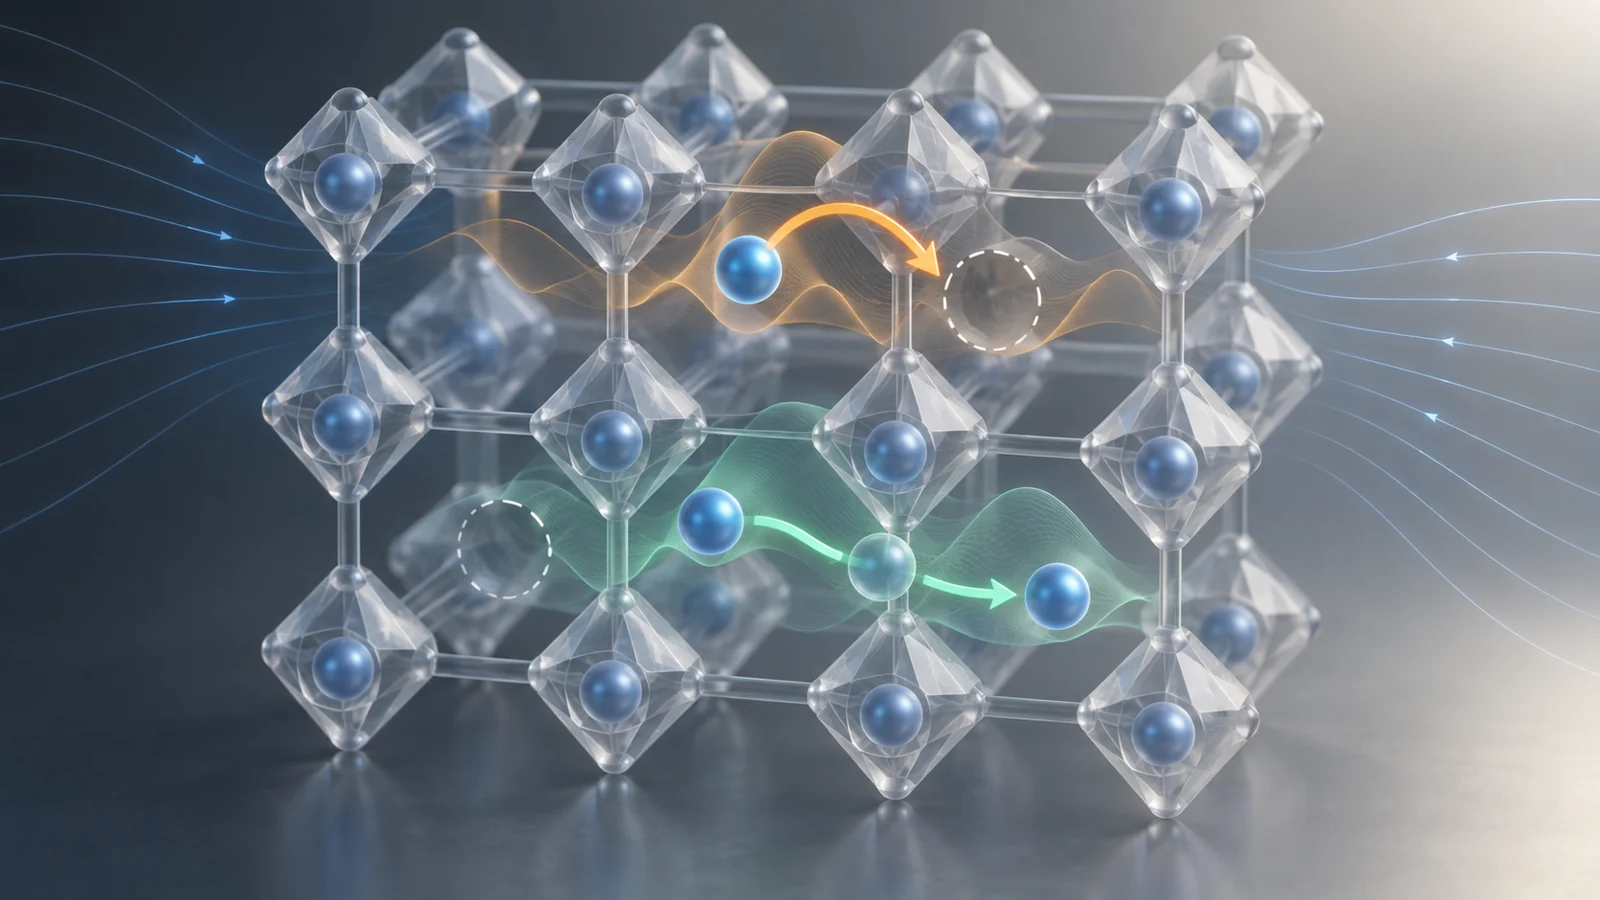
The proposed description begins with ionization reactions that make positive lithium ions available. The model then separates those ions into two transport-related forms.
Some lithium ions hop into neighboring vacancies. In this picture, a vacancy is an available site that can be filled by a nearby ion. Other lithium ions move in a metastable interstitial state rather than following the same vacancy-filling process. The two forms therefore represent different kinds of motion within the same electrolyte description.
That separation also changes how the charged species are represented mathematically. Positive lithium ions are the only moving species in the model. The concentration of negatively charged vacancies is obtained from the governing equations rather than treated as a second population of independently traveling particles. As lithium ions fill some vacancies and occupy other states, the calculated vacancy distribution can change even though the model does not give vacancies their own transport path.
This is why lithium content alone is an incomplete description of transport. Two materials could be assigned the same total lithium concentration while producing different modeled responses if their equilibrium partition between mobile and other states differs, or if the parameters governing the two mechanisms differ. The model’s purpose is to keep those distinctions visible in the equations.
Keeping the energy account intact across scales
The authors also address a mathematical problem that appears when an electrolyte model is meant to connect detailed material behavior with larger-scale calculations. Transport, chemical reactions, electric fields, and charge distributions must fit together without quietly losing the energetic meaning of their interactions.
The formulation gives explicit attention to electromagnetic energy accounting. The paper distinguishes a treatment based on the general Maxwell equations from replacing those equations with electroneutrality alone. That distinction matters because electroneutrality can simplify a calculation, but it does not automatically preserve the same description of electromagnetic energy.
The model’s weak form is constructed to retain a proper energy meaning by using chemical-potential test functions in the mass-balance equations. In practical terms, this gives the mathematical system a disciplined way to connect chemical transport and electromagnetic effects. The authors present the resulting formulation as fully three-dimensional and compatible with linking descriptions at different scales.
Multiscale compatibility here describes the structure of the model, not a completed battery-design workflow. A formulation can be designed so that it is suitable for passing information between scales while still requiring further work before it can reliably represent particular materials, interfaces, or whole cells. The distinction is important: the paper develops a route for calculation, rather than showing that a finished battery simulation has already delivered a better operating result.
Equilibrium and operation ask different questions
The paper examines two related but distinct situations. At equilibrium, the system describes how the species and fields settle into a balanced state. At steady state, the equations describe a sustained operating condition under which quantities may remain stable in time while transport continues.
The governing equations can be solved in closed form for the equilibrium and steady-state cases discussed in the paper. Those solutions make the influence of material parameters easier to inspect. Instead of treating the model as a black box that produces one transport prediction, the analysis shows how changes in its inputs can alter the result.
One input stands out in the reported sensitivity analysis: the fraction of lithium that resides in the mobile state at equilibrium. The authors identify this fraction as the most sensitive parameter within that analysis. That finding gives the model a practical point of focus. If the assumed equilibrium mobile fraction changes, the predicted electrolyte response can change substantially, so estimating that parameter accurately may matter more than refining less influential inputs.
The simulations use a mobile-lithium fraction of 0.64 in the parameter table. That number is a selected model input for the calculations, not a universal property of solid electrolytes. The sensitivity result is likewise tied to the analysis performed in this model; it should not be treated as a ranking that automatically applies to every electrolyte or battery architecture.
A useful calculation still depends on its transport assumptions
The paper’s structure gives the equations a coherent way to represent reactions, chemical potentials, fields, and two lithium-ion mechanisms. But mathematical consistency and physical adequacy are separate questions.
The authors explicitly question whether the classical Nernst–Planck thermodynamic description is appropriate for both transport mechanisms. They note that this form is widely used, while also recognizing that applying it to both the hopping and interstitial descriptions may be a strong assumption. More intricate Maxwell–Stefan free energies are identified as a subject for future work.
That reservation does not erase the model’s contribution. It identifies where the connection between the equations and a real solid electrolyte needs especially careful examination. A model may be internally organized and sensitive to its parameters while still requiring evidence that its chosen transport laws capture the relevant material behavior. Those are different tests: one concerns how the calculation is built, another concerns which inputs control it, and a third concerns how well the description corresponds to the electrolyte being studied.
The result is therefore best read as a more resolved starting point for electrolyte transport modeling. It tells researchers which motions the calculation distinguishes and which parameter deserves close attention, while leaving the adequacy of the shared transport description as an important scientific question.
What this could change
If the two-mechanism description proves suitable for relevant solid electrolytes, and if its key parameters can be measured or constrained with enough accuracy, it could be incorporated into larger battery simulations. Those simulations could then represent electrolyte transport with more information than a single total-lithium concentration provides.
The conditional payoff is better design guidance. Researchers could use the model to compare electrolyte properties with the current demand a cell is intended to support, helping them choose materials whose transport behavior fits the rest of the battery design. That would be a route toward matching electrolyte properties to electrodes and operating requirements before relying on a full-cell result.
As of July 2021, this remains a modeling capability under development. The preprint refines how lithium motion and parameter dependence can be represented, but it does not by itself show that a battery built with those calculations will operate longer or perform better. The next value of the approach depends on whether its assumptions hold for real materials and whether the resulting electrolyte description can be connected credibly to larger, validated cell models.
Key Sources
- Primary source: A two-mechanism and multiscale compatible approach for solid state electrolytes of (Li-ion) batteries
arXiv · Preprint · 2021-07-06