
Since the previous research note · Earlier expectation not tested
Track continuity
Since the previous research note: Could a molecular coating ease solid-state battery manufacturing?
Previous expectation: A water-repelling molecular treatment could give manufacturers more flexibility in handling sulfide electrolytes if it survives realistic processing without harming battery function.
What this study adds: The current simulations identify an isolated lithium cluster within the lithium–Li6PS5Cl interphase and propose an electronic explanation for its formation. This adds a mechanism-focused milestone concerning the assembled material interface, without testing the earlier handling forecast.
What changed in this study
A lithium-metal battery can look straightforward in cross-section: a metal electrode meets an electrolyte, and lithium ions move between the two. In an all-solid-state design, however, the contact itself can become chemically active. A December 17, 2024 arXiv preprint uses atomistic simulations to examine a basic but consequential question: where can unwanted lithium first appear at that boundary?
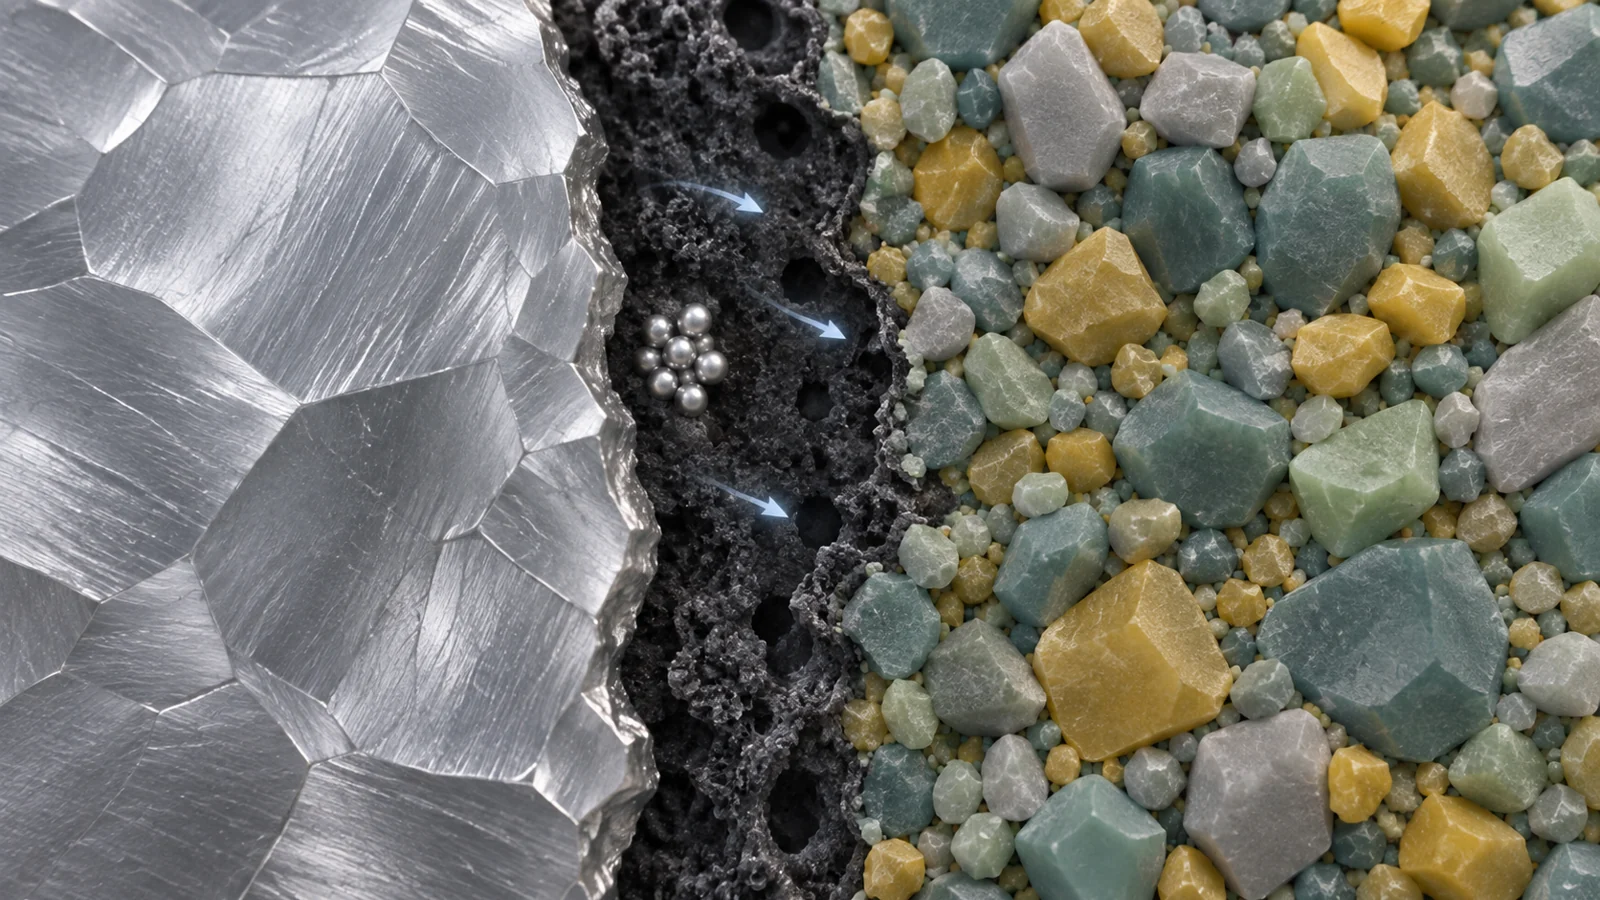
The study reports an isolated lithium cluster forming inside the thin reaction layer between lithium metal and the solid electrolyte Li6PS5Cl. That location is roughly 1 nanometre from the boundary with the metal electrode. The result does not show a complete dendrite or a battery failure, but it shifts attention toward a part of the interface that can be easy to overlook.
The important location is between two familiar regions
The simulated interface has three neighboring regions. One is lithium metal, which acts as the anode. Next comes the solid-electrolyte interphase, or SEI: a reaction layer that develops where the metal and electrolyte meet. Beyond it is Li6PS5Cl, the solid electrolyte through which lithium ions can move.
Before this work, several starting locations could be considered for lithium clustering. A cluster might form directly on the metal surface. It might appear within the bulk solid electrolyte. Or it might develop inside the SEI between them. The simulations identify the third location in the modeled Li–Li6PS5Cl interface.
The spatial detail matters because the cluster is separated from the metal surface rather than simply extending outward from it. If a similar event occurs in an operating cell, a reaction layer could contain a site where lithium ions are converted into metallic lithium before a visible branch reaches out from the electrode. That makes the chemistry and electronic structure of the contact part of the question of how damaging growth begins.
This is a simulated atomic-scale result, not direct imaging of a working battery. Its value is more specific: it provides a possible starting picture for lithium formation at one modeled material interface, with a location precise enough to connect to a proposed chemical mechanism.
How the simulation reached a hard-to-see event
The formation of a lithium cluster is a difficult process for ordinary molecular dynamics to capture. It requires atoms and ions to cross a kinetic barrier, and an unbiased calculation may not encounter that rearrangement during a practical simulation. The researchers therefore combined a learned model of atomic interactions with an enhanced-sampling method.
First-principles calculations based on density functional theory supplied energies and forces for representative structures, including lithium, Li6PS5Cl and their interface. Those data were used in an automatic training workflow to build a deep potential, a machine-learned approximation of the atomic interactions. The trained model made it possible to explore a much larger system than would be practical if every configuration were calculated directly with first-principles methods.
The resulting model was used in molecular-dynamics simulations of a symmetric lithium–Li6PS5Cl–lithium cell containing 4,833 atoms. To make the rare formation event accessible, the researchers used a moving restraint connected to structural coordination variables. In effect, the restraint encouraged the simulation to explore configurations in which lithium atoms became more clustered, while the atomic trajectories supplied the detailed picture of where the cluster emerged.
That procedure is important for interpreting the result. Enhanced sampling is a way to investigate a difficult transition; it is not an ordinary battery clock. The appearance of the cluster in a restrained trajectory cannot be read as the time a real cell would need to develop a dendrite, nor does it provide an unbiased nucleation rate.
A smaller bandgap offers a route for reduction
The proposed explanation begins with lithium-ion reduction. A lithium ion becomes metallic lithium when it gains electrons. For that to happen inside the reaction layer, electrons must be able to reach the region where lithium ions are being reduced.
The authors calculate that the spontaneously formed SEI has a substantially decreased electronic bandgap. A bandgap is the energy separation between occupied electronic states and states that are available for electrons. A smaller separation can make electronic movement through the layer more accessible than it would be in a wider-gap structure.
In this account, the interphase is not merely a passive boundary between the electrode and electrolyte. Its local electronic structure could provide a route for electrons to enter the reaction layer, where they could reduce lithium ions to metallic lithium atoms. That proposed route helps explain why the isolated cluster appears within the SEI rather than directly at the metal surface.
The calculation supports a mechanism, not a direct measurement of electronic conductivity through a working interphase. The distinction matters. A calculated electronic structure can show why electron-enabled lithium formation is chemically plausible in the modeled configuration, while the behavior of a real interface may depend on how that layer forms, changes and interacts with its surroundings.
What the benchmark does and does not settle
The learned interaction model was tested against first-principles calculations on configurations selected for that purpose. The reported root-mean-square errors were 5.45 meV per atom for energy and 93.9 meV per ångström for force. Agreement at that level supports using the model for the sampled configurations and calculated quantities.
It does not turn the entire simulated pathway into an experimentally established account. The benchmark addresses whether the learned model reproduces the reference calculations in the test set. It does not establish that the moving-restraint trajectory follows the natural probability of nucleation, or that the isolated cluster proceeds through the full sequence to a cell-spanning dendrite or short circuit.
The result is also tied to a particular material pairing and modeled interface. The same starting location or electronic explanation may not govern every solid electrolyte, interphase composition or operating condition. Nor does the study, on the evidence considered here, demonstrate a treatment that suppresses dendrites, quantify a lifetime improvement or establish a commercial safety gain.
Those limits leave the central contribution intact. The work connects a specific location—inside the reaction layer, about 1 nanometre from the metal-side boundary—to a specific proposed reason: a local electronic structure that could let electrons participate in lithium formation there. That is a narrower claim than solving dendrite growth, but it is more useful than treating the interface as an undifferentiated contact.
What this could change
If the proposed mechanism also operates in working cells, it could give battery developers a more precise target for interface design. An interphase that limits electron-enabled lithium formation while still allowing lithium-ion transport could reduce the chance that isolated metallic clusters become the starting points for later damage.
Such a strategy would matter because longer-lived lithium-metal solid-state batteries depend on more than the properties of the bulk electrolyte. They also depend on whether the assembled contact remains stable as lithium moves through it. A mechanism-focused understanding of where lithium can first form could therefore support interfaces designed for greater durability and fewer dendrite-related failures.
That benefit remains conditional. The computational result identifies a plausible route at one lithium–Li6PS5Cl interface; it does not demonstrate that an engineered layer can block the route in a functioning battery. The potential payoff is a shift in where developers look for control: not only at the metal surface, but also within the reaction layer that connects the metal to the solid electrolyte.
Key Sources
- Primary source: Observing Li Nucleation at Li Metal-Solid Electrolyte Interface in All-Solid-State Batteries
arXiv · Preprint · 2024-12-17